TL;DR
The FDA says BD and CareFusion have updated a recall notice for certain infusion sets used with the BD Alaris Pump Module 8100, adding that some sets perform worse than previously disclosed. The agency classifies the action as its most serious recall type; facilities are advised to discard listed discontinued sets, review performance warnings and consider alternatives based on clinical needs.
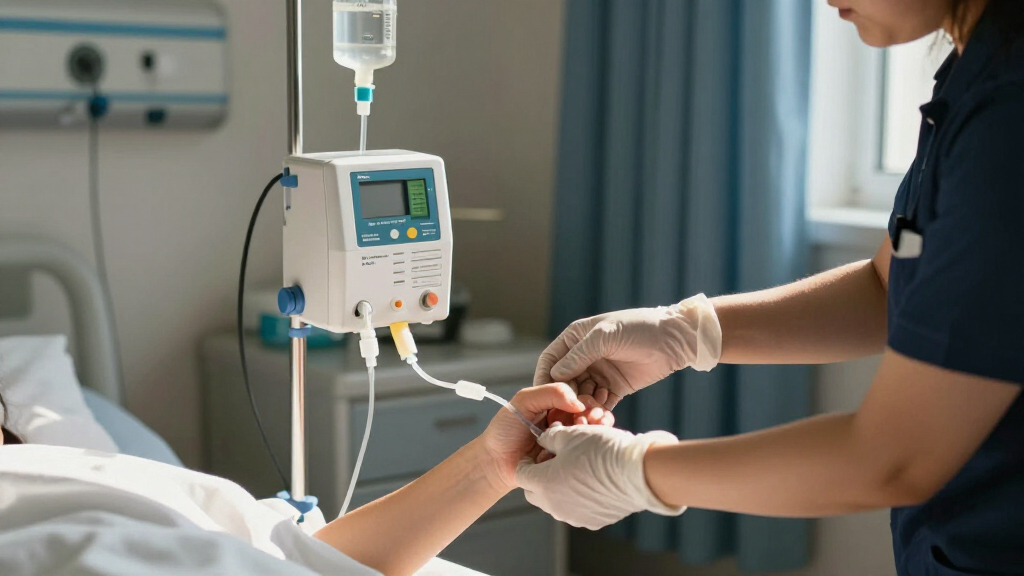
The FDA says BD and its subsidiary CareFusion 303, Inc. have updated guidance for certain infusion sets used with the BD Alaris Pump Module 8100, including instructions to discard sets identified as discontinued. The FDA classifies the action as its most serious type of recall, warning that continued use of affected devices may cause serious injury or death.
In a customer letter dated August 19, 2026, BD and CareFusion described worse-than-previously-disclosed performance under some conditions and updated information about removing and discontinuing certain Alaris Pump Infusion Sets. The FDA says the new performance information replaces material previously communicated in BD field action MMS-25-5311. The devices are used with the Alaris Pump Module model 8100 and Alaris software with Guardrails Suite MX, versions up to 12.6.
BD said testing found that earlier estimates of infusion-set performance were overestimated. Sets with restrictive components or designs — including closed male luer connectors, filters, check valves, Y-sites and microbore tubing — may have larger deviations than previously reported. The company said problems occur most commonly at programmable rates below 1 mL per hour and at or below 5 mL of bolus volume.
The FDA says the potential performance differences include under- or over-infusion, delayed occlusion alarms and inaccurate bolus doses. The agency notes that users may not readily detect these issues, and BD’s letter says the performance data and circumstances of use should guide decisions about set selection and pump use. As of August 19, BD had reported no serious injuries or deaths associated with the issue, according to the FDA.
Risks for Vulnerable Patients
The issue matters because infusion pumps deliver fluids and medicines continuously or intermittently, and an undetected difference between the programmed and delivered amount may affect patient care. The FDA says the possible consequences depend on the circumstances; it identifies neonates, children and critically ill patients as higher-risk groups because they are more likely to receive low-rate or low-volume infusions.
BD cites medicines such as antiarrhythmics and blood-pressure-supporting drugs as examples of treatments that may be delivered at lower rates or volumes. The agency warns that outcomes for vulnerable patients could include serious adverse events or death, but it does not report that such an outcome has occurred in connection with this issue as of the stated date.
The updated information also affects practical decisions at hospitals and other facilities: which sets to remove, how to allocate remaining alternatives, and whether a particular pump-and-set combination remains appropriate for a clinical use. Those decisions must be made with the device warnings and the patient’s care needs in view.
BD Alaris infusion set replacement
As an affiliate, we earn on qualifying purchases.
As an affiliate, we earn on qualifying purchases.
Scope of the Alaris Notice
The affected products are BD Alaris Pump Infusion Sets compatible with Pump Module model 8100. The FDA notice covers use with BD Alaris and Guardrails Suite MX software through version 12.6. It directs readers to the affected-product list and BD’s customer letter for the specific set identifiers; the notice says products listed in the letter’s Attachment C are discontinued.
This update is not only a notice about discontinuing selected sets. It also revises performance information for sets that may remain in use, based on BD’s further testing. That distinction is reflected in the FDA’s recommendations: facilities are told both to remove the identified discontinued products and to review the performance disclosures for decisions about other sets and pump use.
The FDA describes the Alaris System with Guardrails Suite MX as a modular infusion pump and monitoring system used to administer fluids through routes including intravenous, intra-arterial, subcutaneous and epidural administration. The notice concerns the infusion-set performance information and identified products, not a statement that every Alaris pump or set is affected in the same way.
“The previously claimed performance was overestimated.”
— BD, as summarized by the FDA
infusion pump compatible infusion sets
As an affiliate, we earn on qualifying purchases.
As an affiliate, we earn on qualifying purchases.
Unknowns in the Updated Notice
The FDA notice does not provide a count of affected sets or facilities, nor does the supplied material state how many products have been removed. Facilities need to consult the affected-product spreadsheet and BD’s customer letter to identify specific set names and product details.
The notice describes potential deviations and the conditions in which they occur, but it does not quantify the risk for each set, medication or patient. It also does not report serious injuries or deaths as of August 19, 2026; that status does not establish whether less severe events have occurred or whether later reports may emerge.
It remains a clinical and operational question how each facility will handle limited alternative supplies. The FDA and BD provide prioritization guidance, but the notice does not state how much replacement stock is available or when supply constraints, if any, will be resolved.
medical infusion set for critical care
As an affiliate, we earn on qualifying purchases.
As an affiliate, we earn on qualifying purchases.
Facility Actions and Follow-Up
Facilities should identify and discard the infusion sets listed as discontinued in Attachment C of BD’s letter, following their own procedures. If alternatives are limited, BD recommends prioritizing clinically appropriate sets for critical-care, neonatal, infant and pediatric areas. Staff should review the performance disclosures in Attachment B and adjust set selection or pump use where needed.
BD’s guidance says that when performance with an identified set is not clinically appropriate, facilities should use another pump, such as the BD Alaris Syringe Module or an alternative syringe pump. If no syringe pump is available, the notice says recalled sets may be used with caution, enhanced clinical monitoring and adherence to previously communicated warnings. These are manufacturer recommendations described by the FDA; clinicians and facilities must apply them to the relevant circumstances.
The FDA also repeats risk-mitigation steps from the Alaris user manual for versions 12.3 through 12.6. These include keeping the solution container no lower than the pump and no more than 21 inches above it; checking that tubing sits over pressure sensors and is not kinked or collapsed; and following specified precautions for low-rate pressure settings and clearing occlusions. Facilities should consult the full manual and BD’s letter for the complete instructions and product-specific details.
discontinued infusion sets for BD Alaris
As an affiliate, we earn on qualifying purchases.
As an affiliate, we earn on qualifying purchases.
Key Questions
Which infusion sets are covered by the recall notice?
The notice covers BD Alaris Pump Infusion Sets compatible with the BD Alaris Pump Module model 8100. Consult the FDA’s affected-product list and BD’s customer letter for exact product identifiers and the sets designated for discontinuation.
What should a facility do with sets listed as discontinued?
BD and CareFusion advise facilities to identify and discard sets listed in Attachment C of the customer letter, following facility procedures. The FDA also directs users to select a clinically appropriate alternative where available.
What performance problems did BD report?
BD said some sets may have greater differences than previously disclosed, including under- or over-infusion, delayed occlusion alarms and bolus-dose inaccuracies. The company said the deviations occur most commonly at rates below 1 mL per hour and at or below 5 mL of bolus volume.
Have serious injuries or deaths been reported?
According to the FDA, BD had not reported serious injuries or deaths associated with the issue as of August 19, 2026. The agency warns that continued use may cause serious injury or death, particularly for vulnerable patients; its notice does not give a later reporting status.
What if a facility has no alternative pump available?
The FDA’s summary of BD’s guidance says recalled sets may be used if no syringe pumps are available, with caution, enhanced clinical monitoring and adherence to previously communicated warnings. Facilities should review the complete customer letter and device instructions and make decisions appropriate to the clinical situation.
Source: primary